A herbal capsule and the powdered root it was made from are held to different microbial standards, tested at different points, and released against different specifications. Confusing the two is why manufacturers either over-test the raw material or under-specify the finished product.
This article is about the finished product specification — which framework applies, how to set limits that survive an audit, and what release testing should actually consist of.
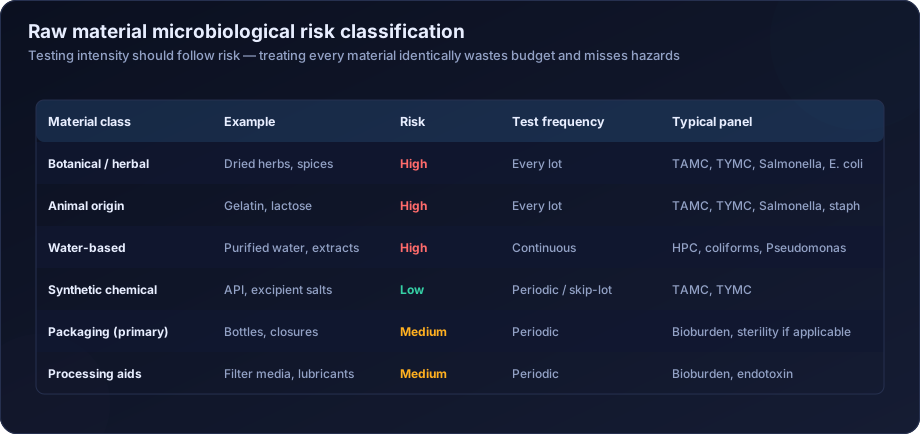
For incoming raw material control — why botanical counts are inherently high, decontamination options and controlling load at source — see botanical raw material microbial contamination. This piece assumes the material is in specification and addresses what happens after.
Which Framework Applies to Your Finished Product
The first question, and the one that determines everything downstream. The same capsule can fall under different frameworks depending on how it is licensed and marketed.
| Product framing | Likely framework in India | Limits basis |
|---|---|---|
| Ayurvedic or herbal medicine | AYUSH / Drugs and Cosmetics Act | AYUSH pharmacopoeial limits |
| Nutraceutical or health supplement | FSSAI | Food safety regulations |
| Pharmaceutical with herbal API | IP / USP / Ph. Eur. | Pharmacopoeial microbial limits |
| Cosmetic with botanical extract | BIS / cosmetic rules | Cosmetic microbiological limits |
| Export product | Destination market rules | Frequently stricter than domestic |
Establish the framework before writing the specification, not after. A specification drafted against pharmacopoeial limits for a product regulated as a food is not wrong so much as arbitrary, and an auditor will ask why those numbers were chosen.
For export products, work to the strictest applicable requirement across your markets. Maintaining separate specifications by destination is possible and creates a segregation burden most manufacturers underestimate.
Route of Administration Changes the Limits
Within any framework, the acceptable microbial load depends on how the product is used, not on what it contains.
Oral solid dosage forms carry the most permissive limits. Products for topical application are tighter. Anything for mucosal, ophthalmic or parenteral use is tighter again, and sterility rather than a limit may apply.
A single botanical extract used across three product formats therefore attracts three different finished-product specifications, even though the input material is identical. This is one of the clearest cases where copying a specification from another product is actively misleading.
Setting a Defensible Specification
Auditors rarely challenge a limit for being too tight. They challenge limits that cannot be justified.
Start from the applicable framework's limit as the ceiling, never as the target.
Look at your own data. If ninety-five percent of batches come in an order of magnitude below the regulatory limit, a specification set at the limit tells you nothing until something has gone badly wrong. An internal alert level set from your own distribution catches drift while it is still correctable.
Distinguish alert from action from specification. The alert level says investigate the trend. The action level says investigate this batch. The specification is the release criterion and the regulatory boundary. Collapsing all three into one number loses the early warning that makes the system useful.
Justify in writing. The rationale — framework, route of administration, historical data, process capability — is what makes the number defensible three years later when the person who set it has moved on.
Review against accumulating data, and be willing to tighten. A specification that has not been revisited in five years of stable production is not being managed.
What Release Testing Should Cover
For a finished botanical product, a proportionate release panel:
| Test | Why it is there |
|---|---|
| Total aerobic microbial count | General hygiene and process control |
| Total yeast and mould count | The indicator that matters most for botanicals |
| Escherichia coli | Faecal contamination indicator |
| Salmonella | Pathogen, absence required |
| Staphylococcus aureus | Where topical or handling risk exists |
| Pseudomonas aeruginosa | Where aqueous or topical |
| Bile-tolerant Gram-negatives | Where the framework requires it |
Do not simply test everything. A panel that includes organisms with no plausible route into your product generates cost and occasional false positives without improving safety. Justify each test by the product, its route and its process.
Method Suitability Is Not Optional
Botanical finished products frequently carry antimicrobial activity of their own — essential oils, tannins, alkaloids, added preservatives. That activity suppresses recovery, and an untreated test can return a clean result on a contaminated product.
Method suitability establishes that your method recovers a known challenge inoculum from your specific product matrix. Without it, a passing result is not evidence of anything.
This must be repeated when the formulation changes, when a supplier or extract source changes, and periodically thereafter. It is among the most common findings in audits of botanical manufacturers, and it is entirely preventable.
Stability and Shelf Life
Microbial specification applies through shelf life, not only at release.
Two failure modes:
Growth during storage, where water activity, packaging or preservation is inadequate. A product released at a low count can drift upward if conditions permit.
Recovery of stressed organisms, where a decontamination step injured rather than killed. Injured cells may not grow at release and may recover during storage, producing a count that rises with time.
The second is the more insidious, and it is the reason a stability programme should include microbial endpoints rather than assay and dissolution alone.
Trending and Investigation
Individual results tell you about a batch. Trends tell you about the process.
Plot counts by product and by batch over time. Look for gradual upward drift, step changes that align with a supplier or process change, and seasonal patterns — botanicals frequently show a monsoon-season signal.
When an out-of-specification result occurs, the investigation should establish whether it is a laboratory error, a genuine product event, or a signal of process drift. Closing it as "laboratory error" without evidence is the most common way a real trend is missed.
Documentation That Supports the Specification
An auditor assessing a finished-product specification asks for three things, and the absence of any one of them is a finding.
The rationale document. Why these limits, referencing the applicable framework, the route of administration, and your own historical distribution. A specification with no written justification is a number somebody chose.
Method suitability records, per product matrix, with the dates and the results. Including repeats after formulation or extract-source changes.
Trend data, showing that the limits are being used as a management tool rather than as a release formality. Alert-level breaches investigated, actions recorded, outcomes documented.
Together these demonstrate that the specification is owned rather than inherited, which is the underlying question every audit of this area is actually asking.
Where Finished-Product Testing Cannot Compensate
A point worth stating plainly, because release testing is sometimes treated as the control rather than as the check.
Sampling cannot find sparse contamination. A pathogen present in a small fraction of a batch will very likely pass a release test. The test confirms the process worked; it does not make an uncontrolled process safe.
Injured organisms may not grow at release. A decontamination step that stressed rather than killed can produce a clean release result and a rising count during storage.
The result arrives after manufacture. By the time a finished-product test fails, the batch exists and the cause is historical.
Which is why finished-product specification sits downstream of raw material control, process hygiene, water quality and environmental monitoring — see botanical raw material microbial contamination. The specification verifies control. It does not create it.
Frequently Asked Questions
Does the specification apply at release or through shelf life? Through shelf life. A product that meets the limit at release and drifts during storage is out of specification, which is why stability programmes should carry microbial endpoints.
Which limits apply to a nutraceutical in India? Usually the FSSAI framework, though a product licensed as an Ayurvedic medicine falls under AYUSH requirements instead. Establish the licensing basis first.
Are herbal limits higher than pharmaceutical limits? Generally the applicable frameworks recognise the inherently higher natural load of botanical material, but the finished-product limit depends on the framework and route of administration rather than on the ingredient.
Can I use one specification across several product formats? No. Route of administration changes the limits, so the same extract in an oral capsule and a topical cream needs different finished-product specifications.
Do I need method suitability for every product? For every product matrix, yes, and repeated when the formulation or extract source changes. Botanical antimicrobial activity makes this essential rather than optional.
Should the specification match the regulatory limit? The regulatory limit is a ceiling. Setting your specification there removes any early warning. Use internal alert and action levels derived from your own data.
What if a batch exceeds the alert level but passes specification? Investigate the trend. That is precisely what an alert level is for, and acting on it is how excursions are prevented rather than managed.
How often should specifications be reviewed? Annually against accumulated data, and whenever the process, supplier or formulation changes.